
What’s New
We are delighted to release our new version of MS Workbook Suite™, Version 2026.
Highlighted Features
- Faster, smarter fragmentation for complex peptide analysis.
- Dynamic labeling for cyclic peptides.
- Save time with fragment transfer.
Administrators are recommended to subscribe to the channels within our support news site found at https://support.revvitysignals.com/hc/en-us/categories/360004446171-Support-News which contains more information about releases and other pertinent product information.
Further Details
Version 2026 adds new features including faster fragmentation, dynamic labeling for cyclic peptides, and faster fragment transfer. There are also improvements in calibration and quantitation, and more.
Data Analysis & Processing
Faster, Smarter Fragmentation for Complex Peptide Analysis
Improved fragmentation algorithms enable faster processing of complex peptide samples, reducing hands-on time, and providing more comprehensive structural characterization, especially for complex cyclic peptides.
- You can now process samples at least twice as fast as previous versions, dramatically reducing analysis time and increasing throughput for high-volume workflows.
- You now have an additional sub-rule to allow for amide bond-reprotonatation, generating more biologically relevant "y-type" ions (Roepstorff-Fohlman-Biemann nomenclature) and improving structural confidence for cyclic peptides.
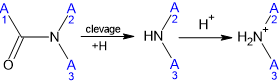
Amide bond reprotontation sub-rule applied to cyclic peptide fragmentation. This enhancement generates additional y-type ions by allowing proton transfer post-cleavage, improving fragment annotation and structural confidence.
- You can now apply fragmentation across all selected components and their corresponding MS1 and MSn components and raw scans with minimal user interaction, freeing analysts to focus on other tasks while processing occurs in the background.
- You can filter the resultant Table of Fragments by ion type (a-, b-, c-, x-, y-, z-, user-labeled, and non-labeled ions), enabling rapid identification of the most relevant fragments and simplifying analysis.
Dynamic Labeling for Cyclic Peptides
Dynamic labeling for cyclic peptides eliminates the need for manual library setup and provides flexible visualization options, accelerating structure characterization and simplifying complex peptide analysis.
- You can now label cyclic peptides dynamically using one-letter and three-letter notations, without adding cyclic structures to the biosequence library to use cyclic peptide ion type labels—saving setup time and reducing library maintenance.
- You can now run AutoAssignment for structures in your preferred view in either expanded (classic) or collapsed (1- or 3-letter notation)—giving you the flexibility to work in the format that best suits your workflow and interpretation needs.
Save Time with Fragment Transfer
- You can now optionally transfer any fragment assignments directly to the query spectrum for the best hit found during an IXCR spectral search into a user-created database—accelerating structure elucidation and reducing manual interpretation time.
Improvements in Quantitation
Enhancements have been made to simplify quantitation and calibration workflows and enhance ease of use, delivering greater flexibility, precision, and efficiency. Using these features, you can now:
- Optionally apply a weighting factor ‘1/x’ to first-degree polynomial calibration curves, enabling more accurate quantitation at lower concentration levels.
- Add any chromatographic parameter to the calibration table, permitting configuration of a single table with all needed information.
- View retention times and compound names as chromatographic descriptors for traces in the LC/UV/MS legend to simplify data interpretation.
- Update the nominal amount directly from the calibration curve, without needing to restart the calibration project.
- Re-access and edit the retention time window for the components after calculation of the calibration curve, saving time when refining calibration parameters.
- Use Summed XICs (molecular and additional ions, that can be satellite, fragment, or adduct ions) for calibration and quantitation, providing more robust quantitation by incorporating multiple related ions.
Store MS3 Component Spectra in Spectrus DB
- You can now store MS3 component spectra in Spectrus DB.
Improved Mass Accuracy
- You can now set up fixed mass accuracy to calculate summed XICs. The following parameters have been introduced:
- XIC accuracy units = 0 (0-Da, 1-ppm)
- XIC accuracy = 0.5 (0.5 Da is default value)
- XIC accuracy (ppm) = 500 (500 ppm is default value)
- Auto calculate summed XIC accuracy = True (by default we use dynamic accuracy)
- You can now set up mass accuracy for Isotope Pattern Calculator in ppm.
Seamless Analysis from a Wider Range of Instrument Types
- You can now change the chromatogram type from xC/UV/MS to HPLC, GC, or CE, enabling seamless analysis of chromatography data from a wider range of instrument types.
Simplified Peak Naming
Save time and reduce errors with enhancements to peak naming when using UV peak matching tools. With these enhancements you can:
- Automatically apply peak names across your entire dataset when using UV peak matching tools, eliminating repetitive manual labeling and ensuring consistent nomenclature.
- Selectively retain previously assigned peak names (except “unknown”) during re-processing, preserving your work and enabling an iterative approach to data analysis without starting from scratch.
Improved Calculation and Display of Isotope Patterns
- You can now set the mass accuracy and calculate the Isotope Pattern based on formulae for manual or Auto Assigned fragments, improving confidence in fragment assignments by providing more accurate isotope pattern matching.
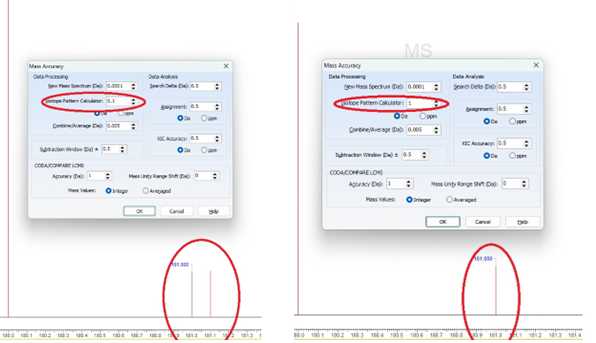
Calculated Isotope Patterns for fragments, left shows Isotope Pattern Calculator set at 0.1 and right shows Isotope Pattern Calculator set at 1.
Improved Visualization of MS/MS Spectra
- You now have the option to “Show Nearest or Component Related Spectra” to display all MS/MS spectra related to the selected component/precursor.
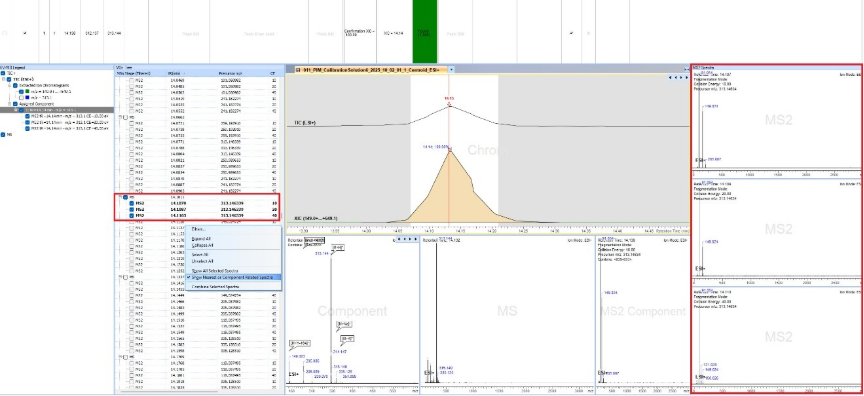
Visualization of three MS/MS Spectra related to the selected component.
Component Spectrum Flexibility
- You can now create component spectra based on the most abundant adduct ion ([M+H]+, [M+NH4]+, [M+K]+, etc.) This is particularly useful when the [M+H]+ ion is weak or exhibits a very low intensity.
Better Handling of Peptide Structures
- Peptides can now be displayed in a simplified form in the Table of Components, making it easier to distinguish and interpret individual structures.
Import Components without Retention Time Assignments
- You can now use existing datafiles to generate target lists (.sdf) for more streamlined batch processing. Components can be imported with or without existing retention times, reducing erroneous assignments and enabling reuse of component files across targeted analysis and other workflows.
Display Combined Retention Time
- You can now display the combined retention time (tR) instead of scan numbers for clearer interpretation of chromatographic data.
Databasing
Create, View, and Edit LC Simulator Files in the Database
- You can now edit and update LC Simulator .lc7 files in Spectrus databases.
- You can now create a Spectrus DB screen form for convenient review of LC Sim Project records.
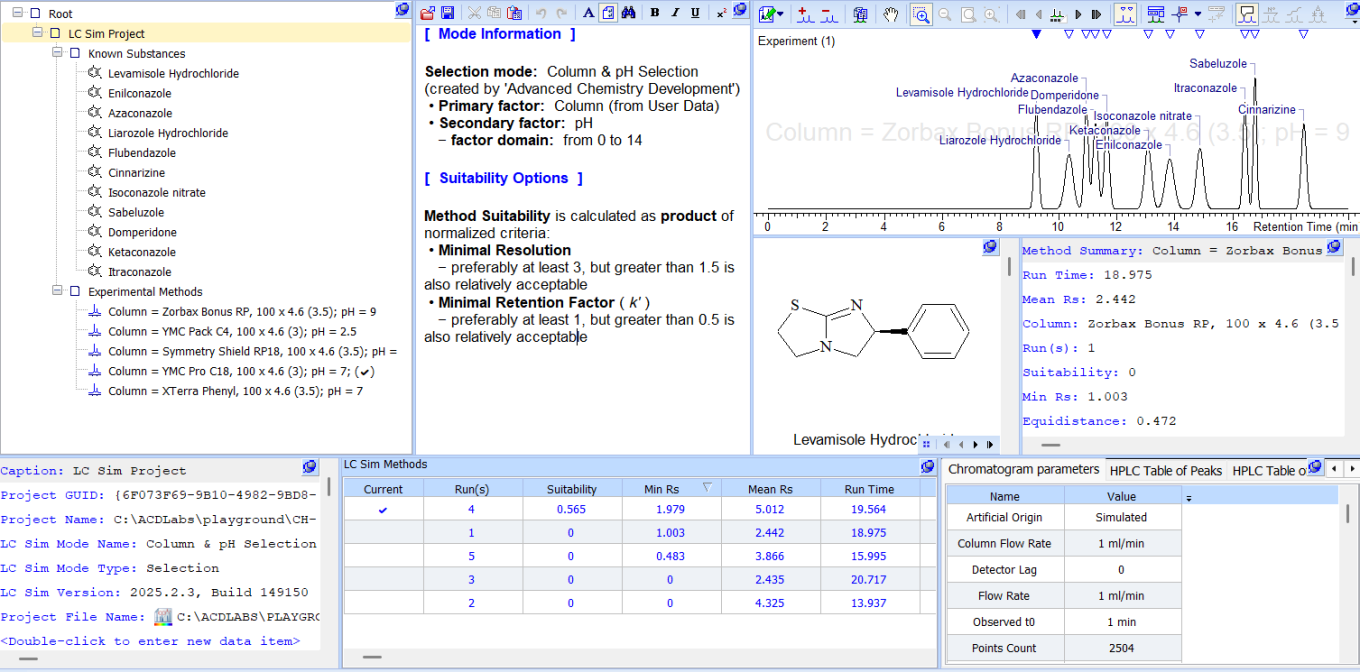
Handle LC Simulator files in database view.
Ease of Use
- Choose to hide the “Custom Screen Form” toolbar for greater screen real-estate.
- You now have a new option to visualize the stereochemical configuration of select stereocenters in database records. Previously you could either display or hide all stereocenters.
Reporting
Improvements to the Reporting Template
- Deconvoluted mass spectra can now be added to the Report by Template along with Raw and Component spectra.
Select Components for Reporting
- You can now choose which components from the Table of Components are included in reports when using the Component/Peak Layout Organizer.
Simplified Peptide Reporting
- You can now report peptides in a simplified form for clearer presentation in reports.
Ease of Use
- You can now populate the “synonym” field in the Table of Components from the same field of the database record (when available).
- You can now navigate directly from the Table of Components to the hit record in the database after running IXCR.
- Settings for IX, IXCR, and ITA are stored along with the individual processed data set/traces.
- “AutoAssignment Options” dialog now displays when the command “Auto Assign All Selected Components” is activated from the Table of Components.
- You can now re-execute a Formulae Generator for selected components (previously you could only run Formulae Generator once for All or Selected components.
- You can now export unassigned components from the Table of Components to an .sdf file.
Data Import & Export
Improved MS & Chrom Data Import
- Improved import filters for Thermo UVPD MSn data.
Include Structure Information in Data Exported in JSON Format
- You can now include structure information in records/databases exported in JSON format via .mol files.
Processing NMR Data
Data Analysis & Processing
Internal Standard qNMR Concentration Calculation Now Included Out of the Box
- Internal standard qNMR calculations are now available in the interface, similar to the external standard workflow (previously available as a script provided upon request).
Conveniently Calculate Similarity Score Between Two 1D Spectra
- You can now calculate the Hit Quality Index (HQI) between two 1D spectra of the same nucleus without the need for the reference spectrum to be in a database. This can provide a quick quantification of the match between the current sample spectrum and that of a standard.
Better Peak Handling for 19F Spectra with Strong Couplings
- You can now automatically pick and assign peaks in 19F spectra of structures containing CHF2 groups more accurately. When the proposed structure contains these groups, the software checks the spectrum for peaks associated with a typical AB spin system for F atoms (two strongly coupled doublets with an evident ‘roof’ effect).
Expanded 1H-31P Coupling Ranges in Auto Assignment
- You can now more accurately assign and verify structures whose spectra contain large 1H-31P coupling constants with increased upper limits for JHP. The new expanded ranges are:
- 3–20 Hz for 2JHP
- 0–20 Hz for 3JHP
- 0–6 Hz for 4JHP
Define Parameters Once for All Replicates in External Standard qNMR
- You can now define parameters including pulse angle, scans, and receiver gain once and have them applied to all selected spectra in external standard qNMR workflows with replicates when they cannot be automatically extracted from the raw data files.
Reporting
Set the Relative Height of “Median” Peak in Report
- You can now better control the “Fit” option for dynamic Y-scaling of the 1D spectra in the reporting templates with the new Ratio parameter that allows you to specify the relative height of the “median” peak in the report.
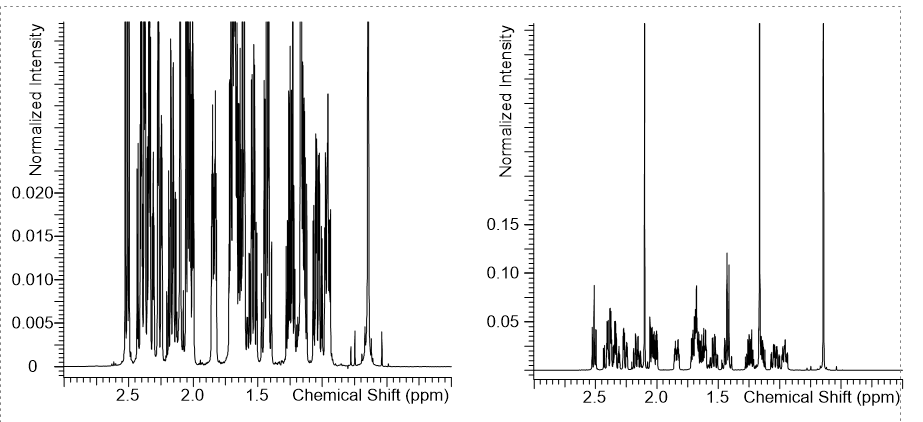
Use the new ratio parameter for dynamic y-scaling of 1D spectra.
Left spectrum has a fit = 90%, right = 10%