
What’s New
We are delighted to release our new version of ChemAnalytical Workbook™, Version 2026.
Highlighted Feature
- Create, view, and edit LC Simulator files in the database.
Administrators are recommended to subscribe to the channels within our support news site found at https://support.revvitysignals.com/hc/en-us/categories/360004446171-Support-News which contains more information about releases and other pertinent product information.
Further Details
Version 2026 adds new features for management of LC Simulator files. There are also improvements to data visualization and data export.
Databasing
Create, View, and Edit LC Simulator Files in the Database
- You can now edit and update LC Simulator .lc7 files in Spectrus databases.
- You can now create a Spectrus DB screen form for convenient review of LC Sim Project records.
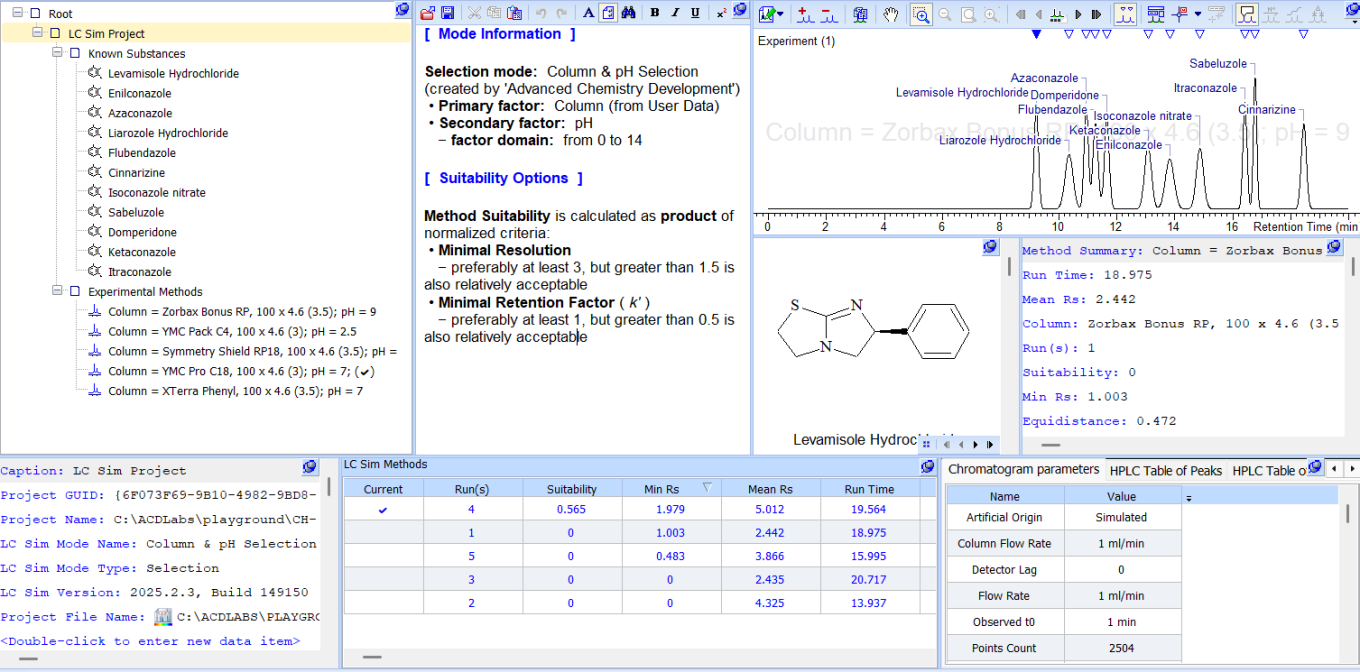
Handle LC Simulator files in database view.
Ease of Use
- Choose to hide the “Custom Screen Form” toolbar for greater screen real-estate.
- You now have a new option to visualize the stereochemical configuration of select stereocenters in database records. Previously you could either display or hide all stereocenters.
Data Import & Export
Import Bio-Sequence Assignments from Any NMR Spectrum
- You can now import assignments for biomolecules from any NMR spectrum in STR format.
Include Structure Information in Data Exported in JSON Format
- You can now include structure information in records/databases exported in JSON format via .mol files.
Improved MS & Chrom Data Import
- Improved import filters for Thermo UVPD MSn data.
Processing MS & Chrom Data
Data Analysis & Processing
Component Spectrum Flexibility
- You can now create component spectra based on the most abundant adduct ion ([M+H]+, [M+NH4]+, [M+K]+, etc.) This is particularly useful when the [M+H]+ ion is weak or exhibits a very low intensity.
Better Handling of Peptide Structures
- Peptides can now be displayed in a simplified form in the Table of Components, making it easier to distinguish and interpret individual structures.
Import Components without Retention Time Assignments
- You can now use existing datafiles to generate target lists (.sdf) for more streamlined batch processing. Components can be imported with or without existing retention times, reducing erroneous assignments and enabling reuse of component files across targeted analysis and other workflows.
Display Combined Retention Time
- You can now display the combined retention time (tR) instead of scan numbers for clearer interpretation of chromatographic data.
Reporting
Select Components for Reporting
- You can now choose which components from the Table of Components are included in reports when using the Component/Peak Layout Organizer.
Simplified Peptide Reporting
- You can now report peptides in a simplified form for clearer presentation in reports.
Processing NMR Data
Data Analysis & Processing
Internal Standard qNMR Concentration Calculation Now Included Out of the Box
- Internal standard qNMR calculations are now available in the interface, similar to the external standard workflow (previously available as a script provided upon request).
Conveniently Calculate Similarity Score Between Two 1D Spectra
- You can now calculate the Hit Quality Index (HQI) between two 1D spectra of the same nucleus without the need for the reference spectrum to be in a database. This can provide a quick quantification of the match between the current sample spectrum and that of a standard.
Better Peak Handling for 19F Spectra with Strong Couplings
- You can now automatically pick and assign peaks in 19F spectra of structures containing CHF2 groups more accurately. When the proposed structure contains these groups, the software checks the spectrum for peaks associated with a typical AB spin system for F atoms (two strongly coupled doublets with an evident ‘roof’ effect).
Expanded 1H-31P Coupling Ranges in Auto Assignment
- You can now more accurately assign and verify structures whose spectra contain large 1H-31P coupling constants with increased upper limits for JHP. The new expanded ranges are:
- 3–20 Hz for 2JHP
- 0–20 Hz for 3JHP
- 0–6 Hz for 4JHP
Define Parameters Once for All Replicates in External Standard qNMR
- You can now define parameters including pulse angle, scans, and receiver gain once and have them applied to all selected spectra in external standard qNMR workflows with replicates when they cannot be automatically extracted from the raw data files.
Reporting
Set the Relative Height of “Median” Peak in Report
- You can now better control the “Fit” option for dynamic Y-scaling of the 1D spectra in the reporting templates with the new Ratio parameter that allows you to specify the relative height of the “median” peak in the report.
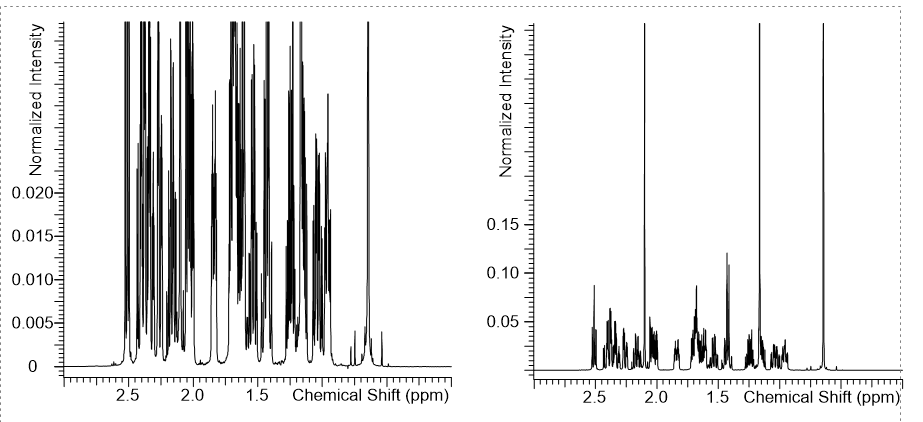
Use the new ratio parameter for dynamic y-scaling of 1D spectra.
Left spectrum has a fit = 90%, right = 10%